Metabolism
Abstract
The hypothesized cellular and molecular mechanisms underlying dystonia are broad and include mutations that perturb Ca2+ signaling, including those affecting voltage gated calcium channels (VGCC). In mice, pharmacological activation of neuronal L-type VGCCs induces dystonia in a dose dependent manner. Here we demonstrate that mice expressing a gain-of-function mutation in the L-type VGCC CaV1.2, associated with Timothy syndrome (TS), exhibit motor dysfunction consistent with dystonia. Although CaV1.2 is broadly expressed throughout peripheral tissues and across the brain, we establish that the dystonia-like behavior is driven by neuronal expression of the mutant calcium channel and observe an associated potential excitatory/inhibitory (E/I) imbalance. Because patients with TS have profound metabolic dysregulation, which is associated with some dystonias, we measured changes in circulating metabolites. The dystonia-like events are sensitive to perturbations in pyruvate metabolism, reminiscent of a subset of dystonias associated with pyruvate dysregulation. Our study provides insight into the potential convergence of previously established causes of dystonia, calcium signaling and metabolic homeostasis.
Authors
Patrick Towers, Hong-Gang Wang, Maiko Matsui, Geoffrey S. Pitt
Abstract
Pneumocystis jirovecii pneumonia (PCP) remains a major cause of life-threatening respiratory failure in people with advanced HIV, yet the biological factors associated with mortality are incompletely understood. We performed integrated transcriptomic, proteomic, and metabolomic profiling of bronchoalveolar lavage fluid from 42 adults with HIV-associated PCP, relating molecular signatures to mortality. We found that fungal burden, measured by sputum immunofluorescence, Pneumocystis qPCR, and 1,3-β-D-glucan, did not differ between survivors and non-survivors or correlate with baseline gas exchange. Instead, non-survivors exhibited a bronchoalveolar immunometabolic failure signature characterised by marked depletion of immunoglobulins and complement, reduced pattern recognition receptors and fibroblast growth factor family proteins, and broad disruption of extracellular matrix organisation. Metabolomic profiling revealed enrichment of arginine-urea cycle and tricarboxylic acid pathways, with coordinated accumulation of arginine, citrulline, and TCA cycle intermediates consistent with nitrosative and mitochondrial stress. An integrated proteo-metabolomic score summarising this state was significantly higher in non-survivors and did not correlate with fungal burden or cytomegalovirus co-infection. In an immunodeficient precision-cut lung slice model, antigen-screened intravenous immunoglobulin restored macrophage-mediated Pneumocystis clearance, providing proof-of-concept for opsonic augmentation. Our findings indicate that mortality in HIV-PCP is defined by profound immunometabolic failure, highlighting humoral depletion and impaired alveolar repair as potential targets for host-directed adjunctive therapy.
Authors
Peter Rossi-Smith, Ayanda Trevor Mnguni, Dora Pungan, Robert J. Samuels, Vaishnavi R. Kumaran, Leena Syed, Trevor Ferris, Kamil Skirlo, Joseph N. Jarvis, Nelesh P. Govender, Graeme Meintjes, Sean Wasserman, Jay K. Kolls, Rachel P.J. Lai
Abstract
The hypothalamic-pituitary-adrenal (HPA) axis integrates central neural processing and responses to stress with endocrine and metabolic responses. We utilized a monoclonal antibody binding corticotrophin-releasing factor (anti-CRF) to achieve durable HPA axis modulation. In mice, anti-CRF treatment decreased plasma corticosterone, food-intake, body weight and fat mass, but preserved lean mass and improved metabolic parameters including insulin sensitization. These body composition effects were observed in middle aged mice on normal chow or high fat diet, aged mice, and Mc4r-/- and ob/ob mice. Single-dose studies demonstrated impacts on weight gain and body composition lasting beyond the period of pharmacological target engagement. Multi-organ transcriptomic and proteomic analyses revealed organ-specific effects of anti-CRF and are consistent with altered glucocorticoid receptor (GR) signaling as a mediator of the observed phenotypes. These studies support anti-CRF-mediated HPA-axis modulation as a metabolic and anti-obesity therapeutic approach distinct from incretins.
Authors
Zachary A. Krumm, Tariful Islam, Caroline M. Watson, Yong Ran, Yona Levites, Hunter S. Futch, Luming Yin, Edgardo Rodriguez-Lebron, Xuefei Liu, Kazi Farhana Afroz, Ariella Goloborodsky, Thomas B. Ladd, Kristy D. Dillon, Danny Ryu, Jennifer L. Bizon, Jada Lewis, Karen A. Scott, Annette de Kloet, Nicholas T. Seyfried, Eric G. Krause, Karen N. McFarland, Todd E. Golde
Abstract
BACKGROUND Dichloroacetate (DCA) is an orally administered structural analog of pyruvate, an endogenous pyruvate dehydrogenase kinase inhibitor.METHODS We conducted a phase III multicenter trial in 34 children with pyruvate dehydrogenase complex deficiency (PDCD). Participants were randomly allocated to 4 months of treatment with DCA or a placebo, followed by a 1-month washout period and crossover to the alternate arm, and could continue into an open-label extension period. DCA dosing was predetermined by pharmacogenomic analysis of GSTZ1, which modulates DCA metabolism. The primary endpoint was the observer-reported outcomes motor domain (ObsROmotor) score. Additional assessments evaluated motor function, plasma lactate levels, and survival.RESULTS Chronic DCA was well tolerated and safe. The primary endpoint, ObsROmotor, was not statistically significantly different between the treatment and placebo groups (P = 0.512). However, longer-term treatment, including the open-label extension, showed a statistically significant treatment effect (P = 0.002), especially in participants with higher baseline motor impairment (ObsROmotor ≥ 8; P = 0.001). DCA decreased plasma lactate –0.48 (0.82) mmol/L (–20%; P = 0.006). Survival of participants was significantly greater than that of a natural history cohort (log-rank P = 0.027).CONCLUSION Longer-term treatment with DCA, dosed based on GSTZ1 haplotype, is safe and was associated with a statistically significant improvement in patient motor function, plasma lactate, and survival.FUNDING NIH (R01FD005407; R42HD089804), University of Florida Department of Medicine, Saol Therapeutics.
Authors
Peter W. Stacpoole, Jose E. Abdenur, Jirair K. Bedoyan, Lorenzo Botto, Gregory M. Enns, Marni J. Falk, Rebecca Ganetzky, Cheryl Garganta, Kevin Glinton, Andrea Gropman, Sharon Hamm, Eugenia Henry, Nicola Longo, Richard Neiberger, Russell P. Saneto, Fernando Scaglia, Sub H. Subramony, Jerry Vockley, Richard E. Wagner

Mitochonic acid 5 alleviates amyotrophic lateral sclerosis phenotypes via mitochondrial augmentation
Abstract
Amyotrophic lateral sclerosis (ALS) is a fatal neurodegenerative disease that urgently requires effective treatment. Mitochondrial dysfunction underlies ALS pathology and represents a potential therapeutic target. Here, we demonstrated the therapeutic potential of mitochonic acid 5 (MA-5), a novel mitochondria-targeted compound that ameliorated ALS phenotypes by enhancing mitochondrial function. In a Drosophila ALS model expressing a mutant human SOD1 (G85R), MA-5 significantly improved locomotor activity, with a trend toward restoration of mitochondrial integrity. In skin fibroblasts derived from ALS patients and motor neurons derived from induced pluripotent stem cells, MA-5 restored ATP production and increased mitochondrial motility. Multiomics analyses suggested that MA-5 modulated mitochondria-linked gene expression and downregulated the glycerophosphate shuttle, contributing to mitochondrial reactive oxygen species production. Transcriptomic analysis identified C7orf31 as a potential marker for monitoring the therapeutic effects of MA-5 and diagnosing ALS subtypes. These findings support MA-5 as a promising therapeutic candidate for ALS and propose C7orf31 as a potential biomarker for treatment monitoring and for disease subtyping.
Authors
Yoshitsugu Oikawa, Yuhan Luo, Naoki Suzuki, Tomoko Kasahara, Yoshiyasu Tongu, Yuki Yoshida, Tsukasa Tominari, Shogo Tanabe, Yoshiko Suto, Hitomi Kashiwagi, Saki Saito, Kensuke Ikeda, Chitose Suzuki, Arata Kuranaga, Tetsuya Akiyama, Satoru Morimoto, Yoshitsugu Aoki, Rieko Muramatsu, Tomoyoshi Soga, Masashi Aoki, Hideyuki Okano, Tetsuhiro Tanaka, Takaaki Abe, Erina Kuranaga, Takafumi Toyohara

Abstract
Foxp3 deficiency causes a profound loss of immune tolerance, unleashing autoreactive T and B cells, lymphoproliferation, cytokine-driven inflammation, and autoantibody production. This autoimmune pathology is fueled by increased glutamine usage, but it remains unresolved whether glutamine is necessary to produce energy or for intermediate metabolite biosynthesis responsible for immunomodulation. Here, we demonstrate that glutamine utilization for biosynthetic pathways supported autoimmune inflammation in the settings of Foxp3 deficiency and dextran sodium sulfate–induced colitis. By employing a model of autoimmunity driven by Treg-specific loss of Foxp3, we showed that this effect is independent of pathogenic Foxp3-deficient Treg reprogramming. Mechanistically, glutamine biosynthetic pathways sustained conventional T cell activation and proinflammatory cytokine production by preventing inosine accumulation and signaling, thus implicating adenosine pathway modulation in autoreactive T cell dysregulation. Conversely, autoreactive B cell activation and autoantibody production relied on glutamine-dependent asparagine availability, which we identified as a targetable vulnerability for autoantibody formation. These findings highlighted glutamine-driven biosynthetic processes as critical drivers of autoimmunity and revealed distinct metabolic vulnerabilities in autoreactive T and B cells that could be targeted for therapeutic intervention.
Authors
Mohammad Adeel Zafar, Charlotte N. Hill Machado, Jyotirmaya Behera, Yuelin Zhong, Xiao Li, Shakchhi Joshi, Yassine El Fazaa, Virginia Camacho, Peter Georgiev, Kiran Kurmi, Marcia Haigis, Louis-Marie Charbonnier
Abstract
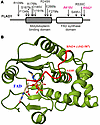
The FLAD1 gene codes for flavin adenine dinucleotide (FAD) synthase. FAD is a cofactor for many redox enzymes involved in vital processes from respiration to signal transduction. In this work, we described a clinical case of 2 siblings carrying compound heterozygous mutations in the FLAD1 gene resulting in the substitutions A418V and R542* at the protein level. The patients demonstrate adrenal insufficiency, which has not previously been associated with FLAD1 protein defects. To verify that adrenal insufficiency is caused by FLAD1 mutations, we created a personalized mouse model carrying the mutations found in the patients. The mutation in the FLAD1 gene, leading to the A418V substitution, appeared viable in the homozygous state, with minimal difference from the WT. The FLAD1 gene mutation leading to the R542* truncation is lethal when homozygous. The mouse model of the compound heterozygous FLAD1A418V/R542* mutations recapitulated the physiological, biochemical, and endocrine manifestations of FLAD1 mutations in patients. The mouse model created demonstrates the causal effect of FLAD1 mutations on the described pathology and potentially paves the way for understanding the disease’s molecular mechanism and developing better therapies.
Authors
Olga A. Averina, Natalia Yu. Kalinchenko, Vitaly A. Ioutsi, Andrey V. Pirogov, Alexander V. Bogachev, Oleg A. Permyakov, Vitaly S. Buev, Ekaterina A. Guseva, Anastasia V. Priymak, Olga A. Bazhanova, Mariia A. Emelianova, Olga O. Grigoryeva, Galina V. Baydakova, Maxim A. Abakumov, Vasily N. Manskikh, Olga A. Dontsova, Petr V. Sergiev, Anatoly N. Tiulpakov

Abstract
Wound healing is a highly dynamic and metabolically demanding process. However, the primary drivers of metabolic alterations involved in this process remain incompletely understood. Here, we employed multiomics profiling of clinical samples to investigate metabolic alterations during wound healing. Our analyses revealed significant activation of the TCA cycle and identified α-ketoglutarate (αKG) as a central regulator orchestrating the reparative phase. Systemic administration of αKG promoted wound closure and re-epithelialization, characterized by enhanced neo-tissue formation with an extended epithelial tongue. Mechanistically, αKG promoted cell proliferation via the cell cycle pathway and enhanced fibroblast-derived TGF-β signaling to induce epithelial-mesenchymal transition–like programs in epithelial cells. To address the spatial metabolic heterogeneity, we developed a transdermal MN platform based on gelatin methacryloyl for localized αKG delivery, further accelerating tissue repair. Collectively, these findings identify αKG as a metabolic driver of wound repair, reveal its dual role in modulating the epithelial-fibroblast microenvironment, and introduce a targeted bioengineering strategy with translational potential for both acute and chronic wound management.
Authors
Yuhan Li, Weimin Lin, Denghao Huang, Yueying Wang, Yimeng Cai, Jie Xiang, Linfeng Liu, Xinxing Shuai, Qi Yin, Shuang Jiang, Malcolm Xing, Yuan Wang, Leixiao Yu, Quan Yuan
Abstract
Gain-of-function (GOF) variants in STAT3 cause a complex disorder characterized by early-onset autoimmunity, lymphoproliferation, recurrent infections, and immune dysregulation. In both primary human and mouse models of STAT3 GOF, CD8+ T cells have been implicated as pathogenic drivers of autoimmunity, though the exact mechanisms remain poorly understood. Here, we found that in patients with STAT3 GOF, CD8+ T cells exist in an activated state. Functional assessment revealed that naive CD8+ T cells have an increased capacity for IFN-γ and TNF-α production, with type I and type II IFN transcriptional signatures. Evaluation of immunoregulatory pathways revealed dysregulation of the purinergic signaling axis in CD8+ T cells: CD39 was increased, whereas downstream purinergic family members, CD73 and the adenosine receptor A2AR, were downregulated, impairing the potential to produce or sense immunosuppressive adenosine. Evaluation of the impact of precision therapy, in the form of JAK inhibition, at a cellular and functional level revealed partial normalization of CD8+ T cell dysregulation in patients, including aberrant cytokine production. Our study suggests that a dysregulated purinergic signaling axis plays a key role in CD8+ T cell dysregulation in STAT3 GOF and may have implications for other rare monogenic immune disorders and common inflammatory disorders.
Authors
Jose S. Campos Duran, Montana S. Knight, Samir U. Sayed, Megan C. Dalalo, Andrea A. Mauracher, Peyton Conrey, Aaron B. Schultz, Ceire A. Hay, Robert B. Lindell, Ilona Neale, Kyle Yeakle, Eric D. Abrams, Erica G. Schmitt, Martin A. Thelin, Christian A. Howard, Sara Bluestein, Christine M. Seroogy, Tamara C. Pozos, Akaluck Thatayatikom, Ingrid S. Lundgren, Amelie Gauthier, Scott W. Canna, Helen C. Su, Michael D. Keller, Ottavia M. Delmonte, Lisa R. Forbes Satter, Steven M. Holland, Jenna R.E. Bergerson, Jennifer W. Leiding, Neil Romberg, Will Bailis, Christopher A. Hunter, Alexandra F. Freeman, Alejandro V. Villarino, Mark S. Anderson, Megan A. Cooper, Tiphanie P. Vogel, Sarah E. Henrickson
Abstract
Inflammatory cytokines reprogram keratinocyte metabolism, but the metabolic pathways that couple immune signals to pathological epidermal growth remain incompletely defined. Here, we identify GLS1-mediated glutaminolysis as a metabolic program preferentially induced in keratinocytes under type 3 inflammatory conditions. Integrated transcriptomic, metabolomic, genetic, and functional analyses showed that IL-17A induced GLS1 expression and glutaminolysis in keratinocytes. Keratinocyte-specific Gls1 deletion reduced the intracellular availability of arginine, proline, and methionine, impaired amino acid-dependent mTORC1 activation, disrupted redox homeostasis, and limited keratinocyte proliferation. Amino acid or antioxidant supplementation partially rescued these defects, whereas rapamycin blocked the amino acid-mediated proliferative rescue. Gls1 deletion did not impair steady-state skin development or homeostasis and did not alter MC903-induced type 2 dermatitis, but it delayed wound re-epithelialization and attenuated IMQ-induced psoriasiform inflammation. Loss of keratinocyte GLS1 also reduced epidermal chemokine expression and the accumulation of neutrophils and IL-17A-producing γδ T cells, revealing a role for glutaminolysis in amplifying epithelial-immune crosstalk. These findings define GLS1-mediated glutaminolysis as a context-specific metabolic checkpoint linking type 3 inflammation to keratinocyte proliferation and cutaneous immune amplification, and support locally or temporally controlled GLS1 inhibition as a potential therapeutic strategy for psoriasis.
Authors
Yongfei Hu, Hai Yu, Kaiyu Liang, Liyan Yuan, Wenjun Zhang, Zhe Zhuang, Suyun Ji, Xichun Xia
No posts were found with this tag.


